R package
Multiple sequence alignments in R, as the interactive JBrowseMSA viewer and as ggplot2 figures.
- In: an alignment (FASTA, Stockholm, Clustal, A3M, or a Biostrings object), a tree (Newick, ape, treeio or ggtree), and the annotations to draw on them, such as domains, genes, highlights, clades or your own column tracks
- Out: an htmlwidget for RStudio, R Markdown, Quarto and Shiny; a ggplot2
figure drawn from the viewer’s own layout, with
msaview_plot(); or the viewer’s own SVG export, withmsaview_save() - Layers: compose with
+, the way ggplot2 and ggtree add a geom
remotes::install_github("GMOD/ggmsaview")Gallery
-
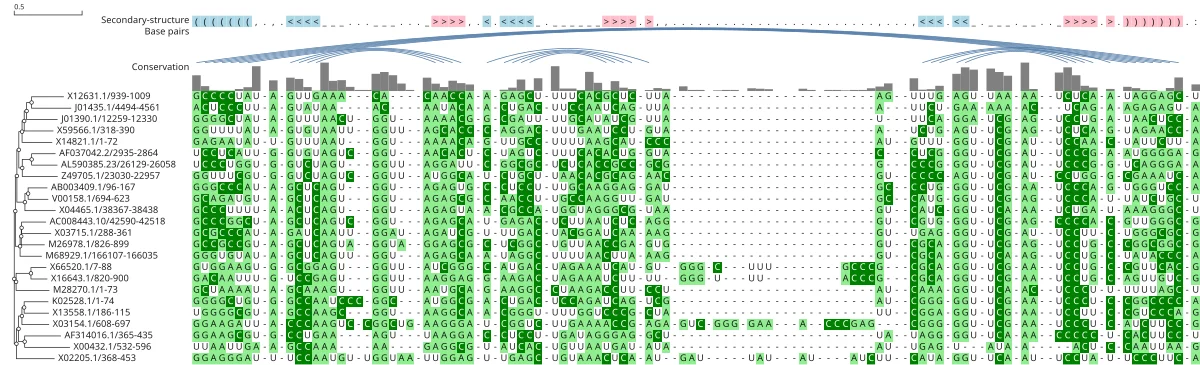
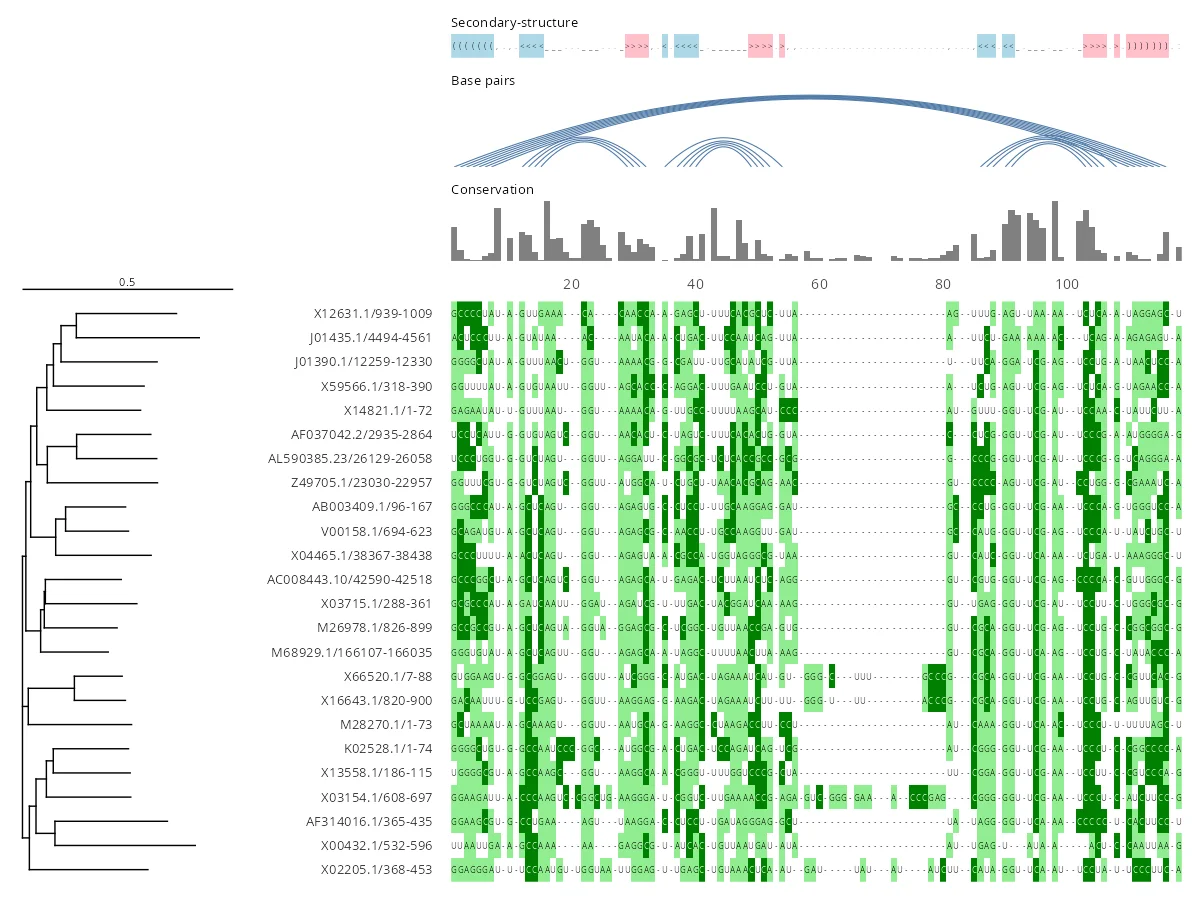
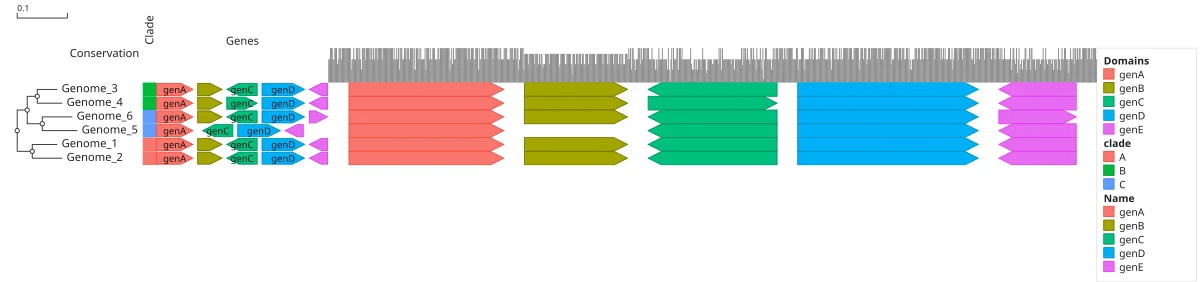
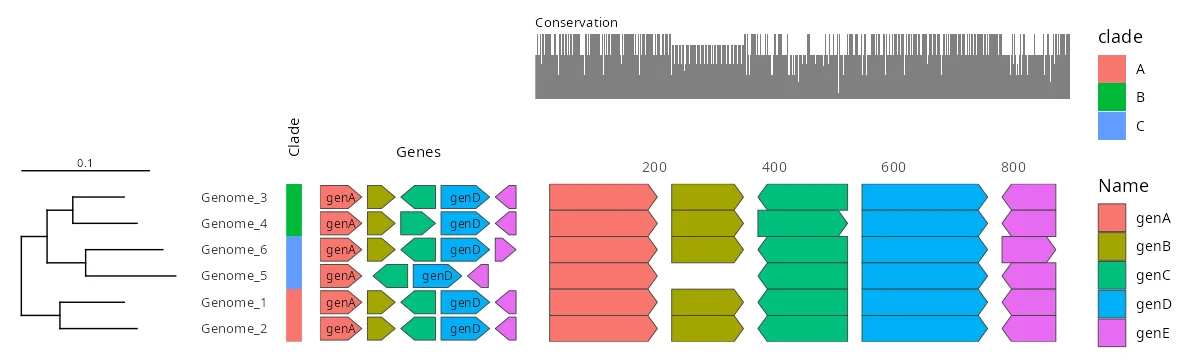
Each figure is drawn twice from data installed with the package: JBrowseMSA’s own SVG export (
msaview_save()) on the left,msaview_plot()on the right. -
Each recipe runs as written after
library(ggmsaview)
e <- function(f) system.file("extdata", f, package = "ggmsaview")
Preproinsulin across nine vertebrates, beside its tree
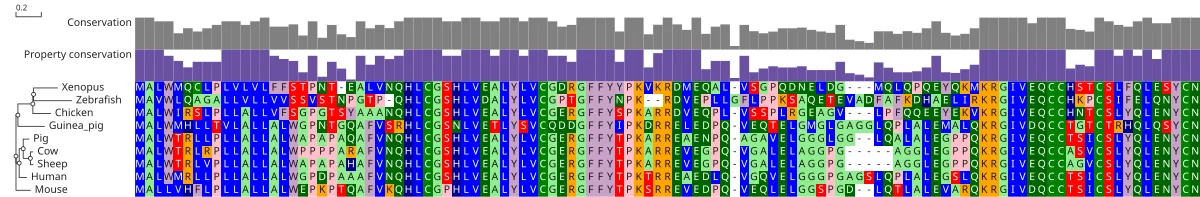
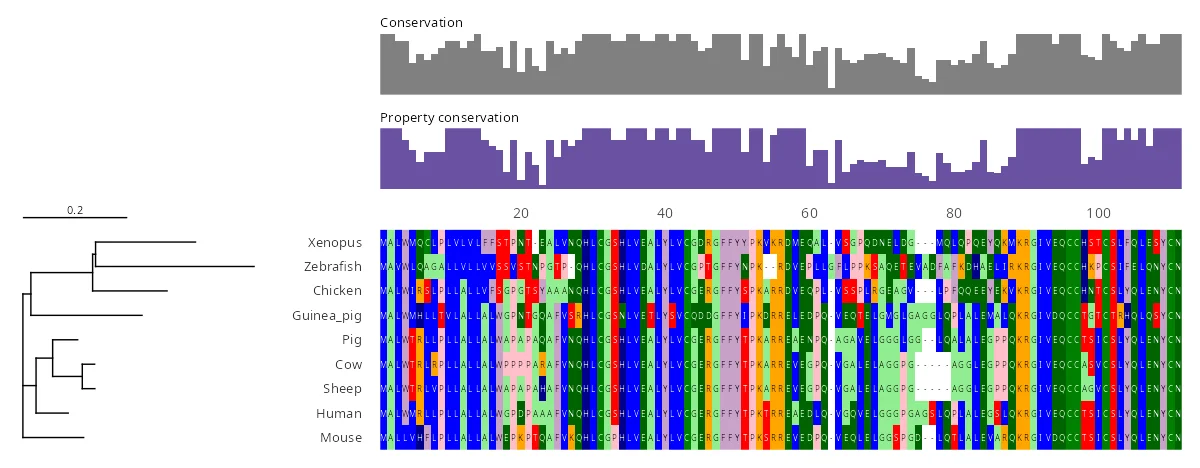
Recipe
w <- msaview(msa = e("insulin.aln"), tree = e("insulin.nh"))
autoplot(w)
msaview_save(w, "figure.svg")
The same alignment drawn as its differences from human
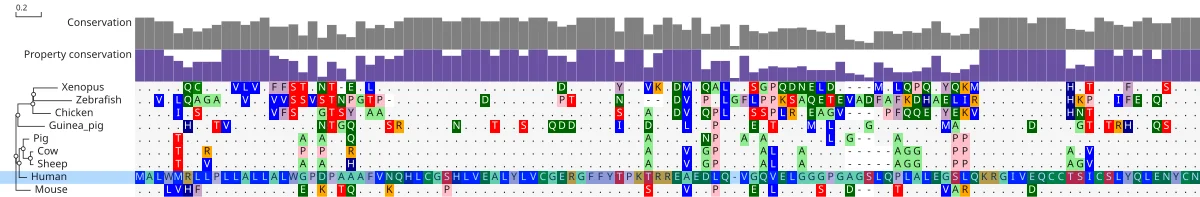
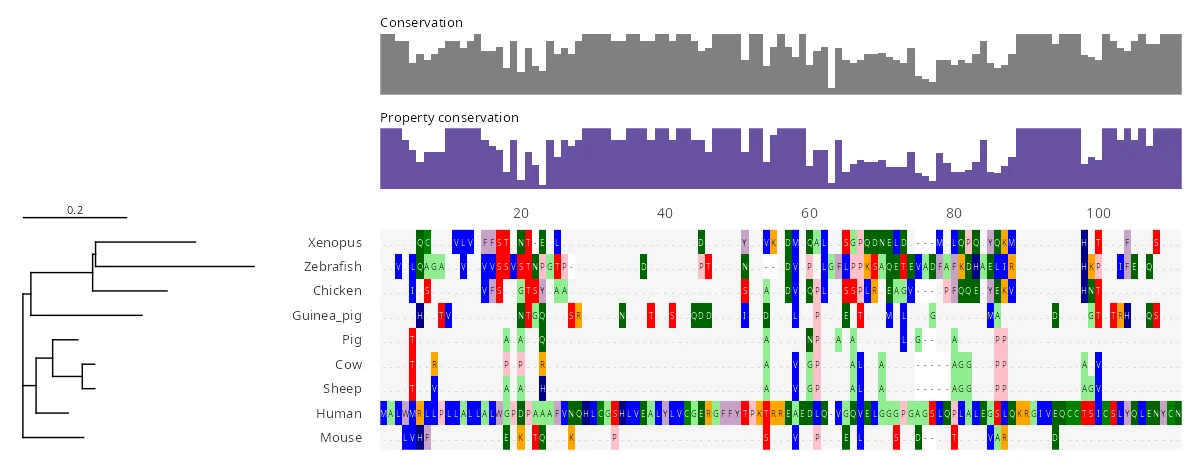
Recipe
w <- msaview(msa = e("insulin.aln"), tree = e("insulin.nh"),
  relative_to = "Human")
autoplot(w)
msaview_save(w, "figure.svg")
Clade brackets over the insulin tree
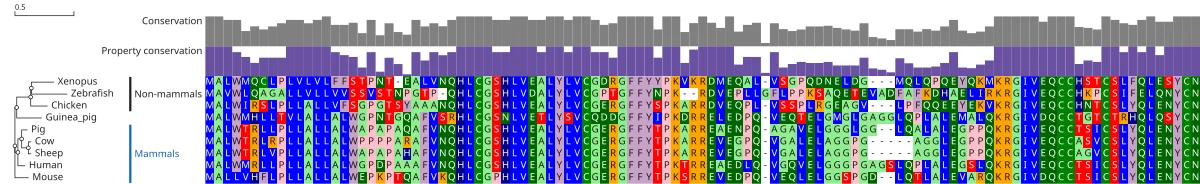
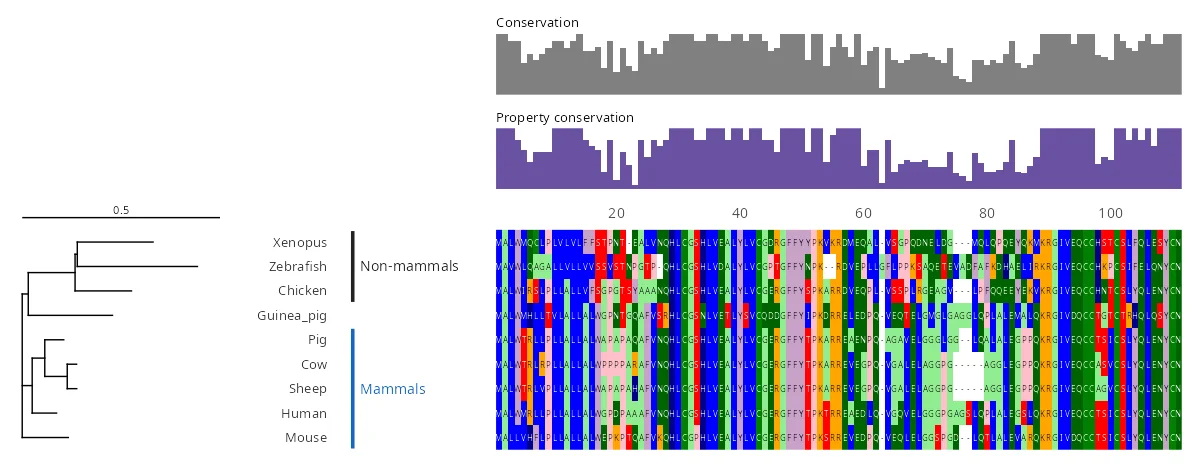
Recipe
w <- msaview(msa = e("insulin.aln"), tree = e("insulin.nh")) +
  geom_msa_clade(range = c("Pig", "Mouse"), tips = 5, mark = "bracket",
    label = "Mammals", color = "#1565c0") +
  geom_msa_clade(range = c("Xenopus", "Chicken"), tips = 3, mark = "bracket",
    label = "Non-mammals")
autoplot(w)
msaview_save(w, "figure.svg")
Ten Src-family kinases with their InterPro domains
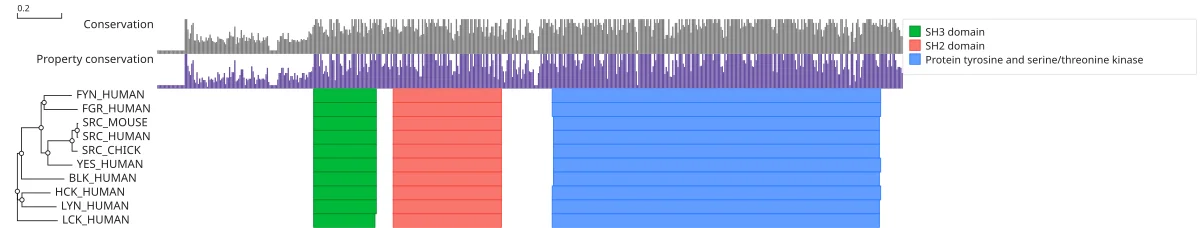
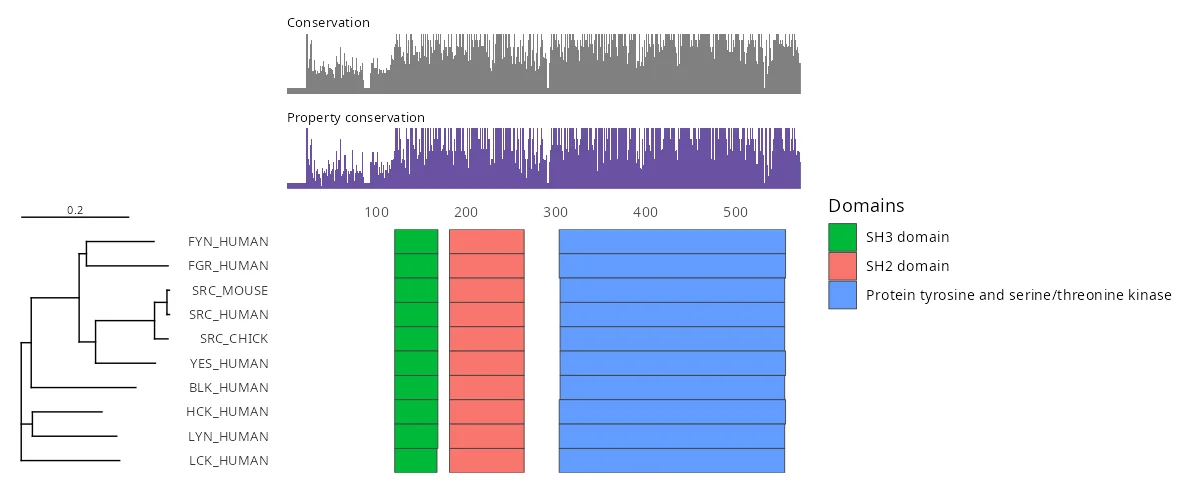
Recipe
w <- msaview(msa = e("kinase.aln"), tree = e("kinase.nh"),
  gff = e("kinase-domains.gff"), col_width = 1.5)
autoplot(w)
msaview_save(w, "figure.svg")
tRNA with its consensus secondary structure
Recipe
w <- msaview(msa = e("trna.stock"))
autoplot(w)
msaview_save(w, "figure.svg")
A gene cluster across six genomes, with each genome’s clade
Recipe
clades <- data.frame(name = paste0("Genome_", 1:6),
  clade = c("A", "A", "B", "B", "C", "C"))
w <- msaview(msa = e("gene-cluster.stock"), gff = e("gene-cluster.gff"),
  col_width = 1) +
  geom_msa_rowdata(clades) +
  geom_msa_strip("clade", header = "Clade") +
  geom_msa_features(x = "position", color = "Name", label = "Name",
    header = "Genes")
autoplot(w)
msaview_save(w, "figure.svg")
Installation
remotes::install_github("GMOD/ggmsaview")The package ships its JavaScript, so it installs with no Node. Only
msaview_save() runs Node, through react-msaview-cli.
For Bioconductor interop:
BiocManager::install(c("Biostrings", "ggtree", "treeio"))Quick start
library(ggmsaview)

widget <- msaview(
  msa = ">human\nMVLSPADKTNVKAAWGKVGAHAGEYGAEALERMFLSFPTTKTYFPHFDLSH\n>mouse\nMVLSGEDKSNIKAAWGKIGGHGAEYGAEALERMFASFPTTKTYFPHFDVSH\n>goat\nMSLTRTERTIILSLWSKISTQADVIGTETLERLFSCYPQAKTYFPHFDLHS",
  tree = "((human:0.1,mouse:0.2):0.05,goat:0.3);"
)
widget
# htmlwidgets::saveWidget(widget, "alignment.html")  # self-contained HTML
# msaview_plot(widget)                                # ggplot2 figure, see below
ggplot2 figures
msaview_plot() draws the same widget with ggplot2, so ggsave(), themes and
patchwork apply:
- The viewer’s own model runs in an embedded QuickJS engine (the
QuickJSR package), so collapsed
clades, hidden gappy columns, color schemes, domains, highlights and
relative_toall come out as they do on screen. No Node or browser runs - The figure draws the viewer’s row panels (strips and gene arrows), its column
tracks (conservation, the sequence logo, arcs, Stockholm text rows and your
own
geom_msa_track()s) - Domains, gene arrows and strips map
fillto their category through ascale_fill_manual()seeded with the viewer’s colors, so ascale_fill_*()you add replaces it, and patchwork collects the legends msaview_layout()returns the tables behind the figure (rows, letter colors, tree segments, feature shapes, row panels, highlights, clades, column tracks) for your own layers- A widget knitted into a PDF or Word document from R Markdown or Quarto prints as this figure
library(ggplot2)
p <- msaview_plot(widget, tracks = c("conservation", "sequence-logo"))
ggsave("alignment.pdf", p, width = 8, height = 2)
SVG from the viewer’s own renderer
msaview_save() writes the SVG the viewer exports, or a PNG or PDF of it
through the rsvg package:
- It needs Node.js, and runs
npx react-msaview-cliunlessoptions(msaview.cli = ...)names another copy - No browser runs, so it works in a batch job or on a server
widthsets the view’s pixel width; the whole alignment is drawn whatever its length- The CLI refuses an alignment of more than 500,000 residue letters, since each
letter is an SVG element; a
col_widthbelow 5 or arow_heightbelow 8 draws it with no letters
msaview_save(widget, "alignment.svg")
msaview_save(widget, "alignment.png", scale = 3)
Examples
A protease family in R is the long version: fourteen UniProt accessions cut to their peptidase S1 domain, aligned with DECIPHER, and drawn with a BLOSUM62 track, disulfide arcs and a band on each catalytic residue.
Layers, composed with +
Every argument of msaview() is also a layer, added the way ggplot2 and ggtree
add a geom. Each layer reads a data frame, so a table of variants or per-column
numbers goes in as it already sits in your session.
library(ggmsaview)

variants <- data.frame(
  row   = c("Human", "Human"),
  start = c(248, 273),
  end   = c(248, 273),
  label = c("R248Q", "R273H")
)

msaview(msa = "p53.aln", tree = "p53.nh") +
  geom_msa_domains("p53-domains.gff") +
  geom_msa_highlight(variants) +
  geom_msa_track(data.frame(value = conservation), name = "Conservation") +
  scale_residue_color("clustal", encoding = "color") +
  stat_msa_diff("Human") +
  coord_msa(100, 288, row = "Human") +
theme_msa("dark", col_width = 14)| Layer | What it adds |
|---|---|
geom_msa_domains() | a GFF3 file, URL, text or data frame of annotations |
geom_msa_highlight() | a data frame of spans, or one span from its arguments |
geom_msa_track() | a track of your numbers, text or arcs |
geom_msa_clade() | a rectangle, bracket, collapse or rotation of a clade |
geom_msa_structure() | which structure residue each row residue is |
geom_msa_rowdata() | a data frame of extra fields per row |
geom_msa_strip() | a color strip beside the tree from one of those fields |
geom_msa_features() | the GFF’s spans per row in a panel beside the tree |
scale_row_color() | colors a tip label or a row tint by one of those fields |
scale_residue_color() | the color scheme, and the channel it paints |
stat_msa_diff() | draws every row as its differences from one row |
coord_msa() | the span the viewer opens on |
geom_msa_selection() | a block of columns and rows, selected |
coord_tree() | the tree’s child order (ladderize) and its root |
theme_msa() | cell size, tree gutter, toolbar, light or dark |
geom_msa_highlight(), geom_msa_clade(), geom_msa_track(),
geom_msa_structure(), geom_msa_strip(), geom_msa_features() and
scale_row_color() accumulate, so calling one twice adds two. The others
replace what an earlier layer set.
A layer builds the same props the matching msaview() argument does, so the two
styles mix freely and produce the same viewer. msa and tree are the viewer
itself and stay arguments; every other argument has a layer, which
test-layer-coverage.R checks.
Named character vector
seqs <- c(
  human = "MVLSPADKTNVKAAWGKVGAHAGEYGAEALERMFLSFPTTKTYFPHFDLSH",
  mouse = "MVLSGEDKSNIKAAWGKIGGHGAEYGAEALERMFASFPTTKTYFPHFDVSH",
  goat  = "MSLTRTERTIILSLWSKISTQADVIGTETLERLFSCYPQAKTYFPHFDLHS"
)
msaview(msa = seqs, tree = "((human:0.1,mouse:0.2):0.05,goat:0.3);")
From files
msaview(msa = "alignment.stock")
msaview(msa = "alignment.fa", tree = "tree.nwk")
With ape
library(ape)

tree <- rtree(15)
seqs <- setNames(
  replicate(15, paste0(sample(c("A","C","G","T"), 300, TRUE), collapse = "")),
  tree$tip.label
)
msaview(msa = seqs, tree = tree, color_scheme = "nucleotide")
Cladogram (no branch lengths)
msaview(msa = seqs, tree = tree, show_branch_len = FALSE)With Biostrings
library(Biostrings)

# DNAStringSet
dna <- DNAStringSet(c(
  seq1 = "ATGCGATCGATCGATCG--ATCG",
  seq2 = "ATGCGATCGATCGATCGATCGATCG",
  seq3 = "ATGCG--CGATCGATCGATCGATCG",
  seq4 = "ATGCGATCGATCGATCG--ATCG"
))
msaview(msa = dna, color_scheme = "nucleotide")

# AAStringSet
aa <- AAStringSet(c(
  human = "MVLSPADKTNVKAAWGKVGAHAGEYGAEALERMFLSFPTTKTYFPHFDLSH",
  mouse = "MVLSGEDKSNIKAAWGKIGGHGAEYGAEALERMFASFPTTKTYFPHFDVSH",
  goat  = "MSLTRTERTIILSLWSKISTQADVIGTETLERLFSCYPQAKTYFPHFDLHS"
))
msaview(msa = aa, color_scheme = "clustal")

# DNAMultipleAlignment
# Biostrings rejects a MultipleAlignment whose rows differ in length
aln <- DNAMultipleAlignment(c(
  seq1 = "ATGCGATCGATCGATCG--ATCG",
  seq2 = "ATGCGATCGATCGATCGATCGAC",
  seq3 = "ATGCG--CGATCGATCGATCGAC"
))
msaview(msa = aln)
After running the msa package
library(msa)
library(Biostrings)

unaligned <- readAAStringSet("proteins.fasta")
aligned <- msa(unaligned, method = "ClustalOmega")
msaview(msa = as(aligned, "AAStringSet"))
Domain annotations
Overlay InterProScan-style protein domains as boxes on the alignment. The gff
argument accepts a file path, a GFF3 string, or a data frame. This pairs with
the CLI, which writes domains as GFF3:
react-msaview-cli interpro accessions.tsv -o domains.gffseqs <- c(
GPCR_human = "MNGTEGPNFYVPFSNATGVVRSPFEYPQYYLAEPWQFSMLAAYMFLLIVLGFPINFLTLYVTVQHKKLR",
GPCR_mouse = "MNGTEGPNFYVPFSNKTGVVRSPFEYPQYYLAEPWQFSMLAAYMFLLIMLGFPINFLTLYVTVQHKKLR",
GPCR_bovine = "MNGTEGPNFYVPFSNATGVVRSPFEYPQYYLAEPWQFSMLAAYMFLLIVLGFPINFLTLYVTVQHKKLR"
)
# from a CLI-generated GFF file
msaview(msa = seqs, gff = "domains.gff")
# or from a data frame: row (or seqname), start, end, and any fields,
# such as a name, a description, a strand or a column an encoding reads
domains <- data.frame(
seqname = names(seqs),
start = 6,
end = 62,
name = "PF00001",
description = "7tm receptor (rhodopsin family)"
)
msaview(msa = seqs, gff = domains, color_scheme = "clustalx_protein_dynamic")
Labeled highlights
Mark a residue, a column range, or a set of rows, with a label saved in the
widget. Coordinates are 1-based and inclusive; row makes start and end
residues of that sequence, projected through the alignment’s gaps.
msaview(
  msa = seqs,
  highlights = list(
    list(row = "GPCR_human", start = 20, end = 26, label = "ligand pocket"),
    list(rows = c("GPCR_mouse"), label = "knockout", color = "rgba(0,120,255,0.2)")
  )
)
A track from your own numbers
column_tracks draws values you compute per column, or per residue of one
sequence, as a track above the alignment. The viewer scales bars by max
(default 1). When row names a sequence, the viewer places each value on that
sequence’s residues and skips the columns where the sequence has a gap.
hydropathy <- c(I = 4.5, V = 4.2, L = 3.8, F = 2.8, C = 2.5, M = 1.9, A = 1.8,
                G = -0.4, T = -0.7, S = -0.8, W = -0.9, Y = -1.3, P = -1.6,
                H = -3.2, E = -3.5, Q = -3.5, D = -3.5, N = -3.5, K = -3.9,
                R = -4.5)
residues <- strsplit(gsub("-", "", seqs[["GPCR_human"]]), "")[[1]]

msaview(
  msa = seqs,
  column_tracks = list(
    list(id = "kd", name = "Hydropathy (human)", kind = "bar",
         values = hydropathy[residues] + 4.5,
         scales = list(y = list(domainMax = 9)), row = "GPCR_human",
         color = "#6a51a3")
  )
)
A kind = "text" track takes data, one character per column, and an optional
encoding = list(color = list(domain =, range =)), a color per character. Every
field is listed in the layers reference.
With ggtree
Pass a ggtree plot object directly as the tree argument, and msaview reads
the tree out of it.
library(ggtree)
library(ape)

tree <- rtree(20)
p <- ggtree(tree) + geom_tiplab()

seqs <- setNames(
  replicate(20, paste0(sample(c("A","C","G","T"), 200, TRUE), collapse = "")),
  tree$tip.label
)

# pass the ggtree plot object directly
msaview(msa = seqs, tree = p, color_scheme = "nucleotide")
ggtree with annotations
Annotated ggtree plots work too. msaview takes only the tree; the colors,
labels and metadata stay in the ggtree plot.
metadata <- data.frame(
  label = tree$tip.label,
  group = sample(c("A", "B", "C"), 20, replace = TRUE)
)
p2 <- ggtree(tree) %<+% metadata +
  geom_tiplab(aes(color = group)) +
  theme(legend.position = "right")

msaview(msa = seqs, tree = p2)
With treeio
library(treeio)

# treedata objects from read.beast, read.raxml, etc.
beast_tree <- read.beast("beast_output.tree")
msaview(tree = beast_tree)

# or convert from phylo
td <- as.treedata(tree)
msaview(msa = seqs, tree = td)
In Shiny
The widget sets three inputs named after its output id. input$<id>_click holds
the cell a click pinned: column is the 1-based column of the file, row the
row name, residue the 1-based position in that row’s sequence (absent on a
gap), and letter the character. It is NULL after a click clears it.
input$<id>_viewport holds the columns on screen as startColumn and
endColumn. input$<id>_selection holds the block a shift-drag selects:
start and end columns of the file, and rows, the selected row names,
absent when the block spans every row. It is NULL after Escape clears it. A
re-render that keeps the same alignment keeps the reader’s scroll and zoom, so
changing color_scheme below restyles the view in place.
library(shiny)
library(ggmsaview)

ui <- fluidPage(
  titlePanel("MSA Viewer"),
  sidebarLayout(
    sidebarPanel(
      fileInput("msa_file", "Upload alignment",
                accept = c(".fa", ".fasta", ".stock", ".sto", ".aln")),
      fileInput("tree_file", "Upload tree (optional)",
                accept = c(".nwk", ".nh", ".newick")),
      selectInput("color_scheme", "Color scheme",
                  choices = c("maeditor", "clustal", "lesk", "cinema",
                              "flower", "nucleotide", "jbrowse_dna",
                              "clustalx_protein_dynamic",
                              "percent_identity_dynamic"),
                  selected = "maeditor"),
      actionButton("example_btn", "Load example data")
    ),
    mainPanel(
      msaviewOutput("msa_viewer", height = "600px"),
      verbatimTextOutput("clicked")
    )
  )
)

server <- function(input, output, session) {
  msa_data <- reactiveVal(NULL)
  tree_data <- reactiveVal(NULL)

  observeEvent(input$msa_file, {
    msa_data(input$msa_file$datapath)
  })

  observeEvent(input$tree_file, {
    tree_data(input$tree_file$datapath)
  })

  observeEvent(input$example_btn, {
    msa_data(paste0(
      ">human\nMVLSPADKTNVKAAWGKVGAHAGEYGAEALERMFLSFPTTKTYFPHFDLSH\n",
      ">mouse\nMVLSGEDKSNIKAAWGKIGGHGAEYGAEALERMFASFPTTKTYFPHFDVSH\n",
      ">goat\nMSLTRTERTIILSLWSKISTQADVIGTETLERLFSCYPQAKTYFPHFDLHS"
    ))
    tree_data("((human:0.1,mouse:0.2):0.05,goat:0.3);")
  })

  output$msa_viewer <- renderMsaview({
    req(msa_data())
    msaview(
      msa = msa_data(),
      tree = tree_data(),
      color_scheme = input$color_scheme
    )
  })

  output$clicked <- renderPrint(input$msa_viewer_click)
}

shinyApp(ui, server)
Supported inputs
| Parameter | Accepted types |
|---|---|
msa | File path, FASTA/Stockholm/Clustal string, named character vector, DNAStringSet, AAStringSet, RNAStringSet, DNAMultipleAlignment, AAMultipleAlignment |
tree | File path, Newick string, ape::phylo, treeio::treedata, ggtree plot object |
Color schemes
Protein: maeditor, clustal, lesk, cinema, flower, clustalx_protein,
jalview_taylor, jalview_zappo, jalview_hydrophobicity, jalview_buried,
jalview_prophelix, jalview_propstrand, jalview_propturn
Nucleotide: nucleotide, jbrowse_dna, rainbow_dna, clustalx_dna
Dynamic (computed per-column): clustalx_protein_dynamic,
percent_identity_dynamic
A named vector of colors keyed by residue letter colors those letters, in either case, and leaves every other letter uncolored:
msaview(msa = aa, color_scheme = c(K = "#1f77b4", R = "#1f77b4", D = "#d62728"))